Research question
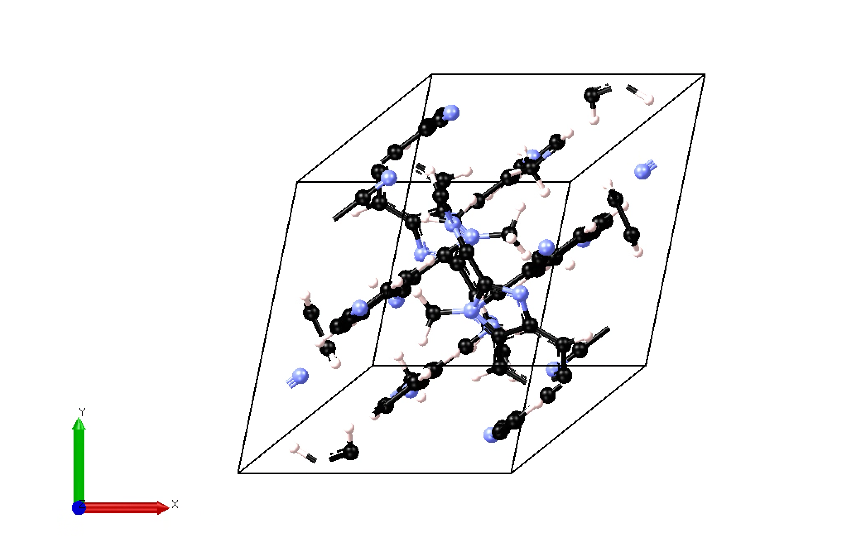
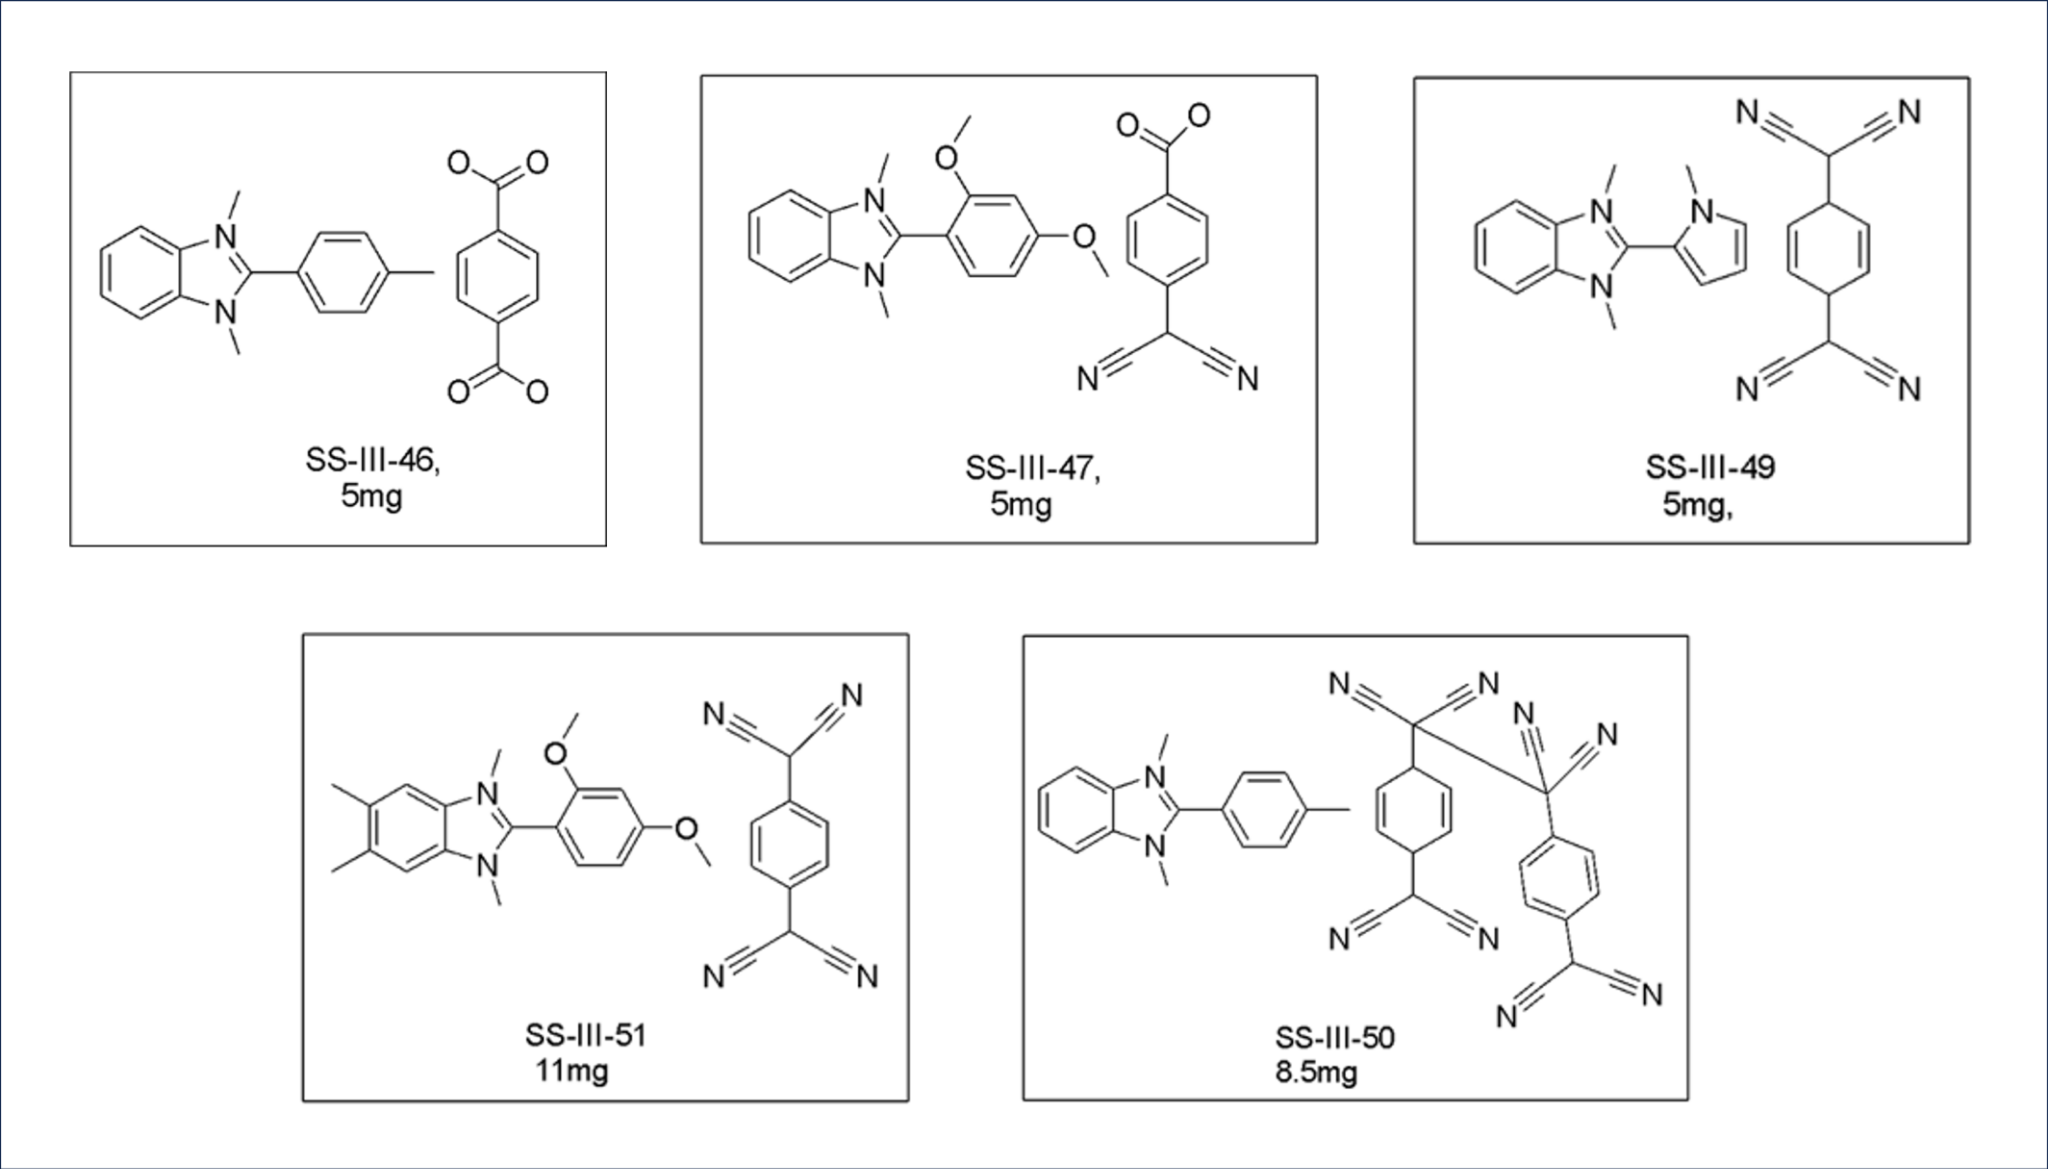
Organic cocrystals combine electron donors and acceptors in a repeating structure. Small changes in the molecules or how they stack can change their electronic properties. Our study examined newly synthesized cocrystals based on a benzimidazole-hydride (BIH) donor and TCNQ derivatives as acceptors.
I investigated their electronic structure with computational calculations on NYU's high-performance computing resources. The work used crystal structures determined by X-ray diffraction by collaborators and compared isolated donor–acceptor pairs with their periodic crystal structures.
Computational approach
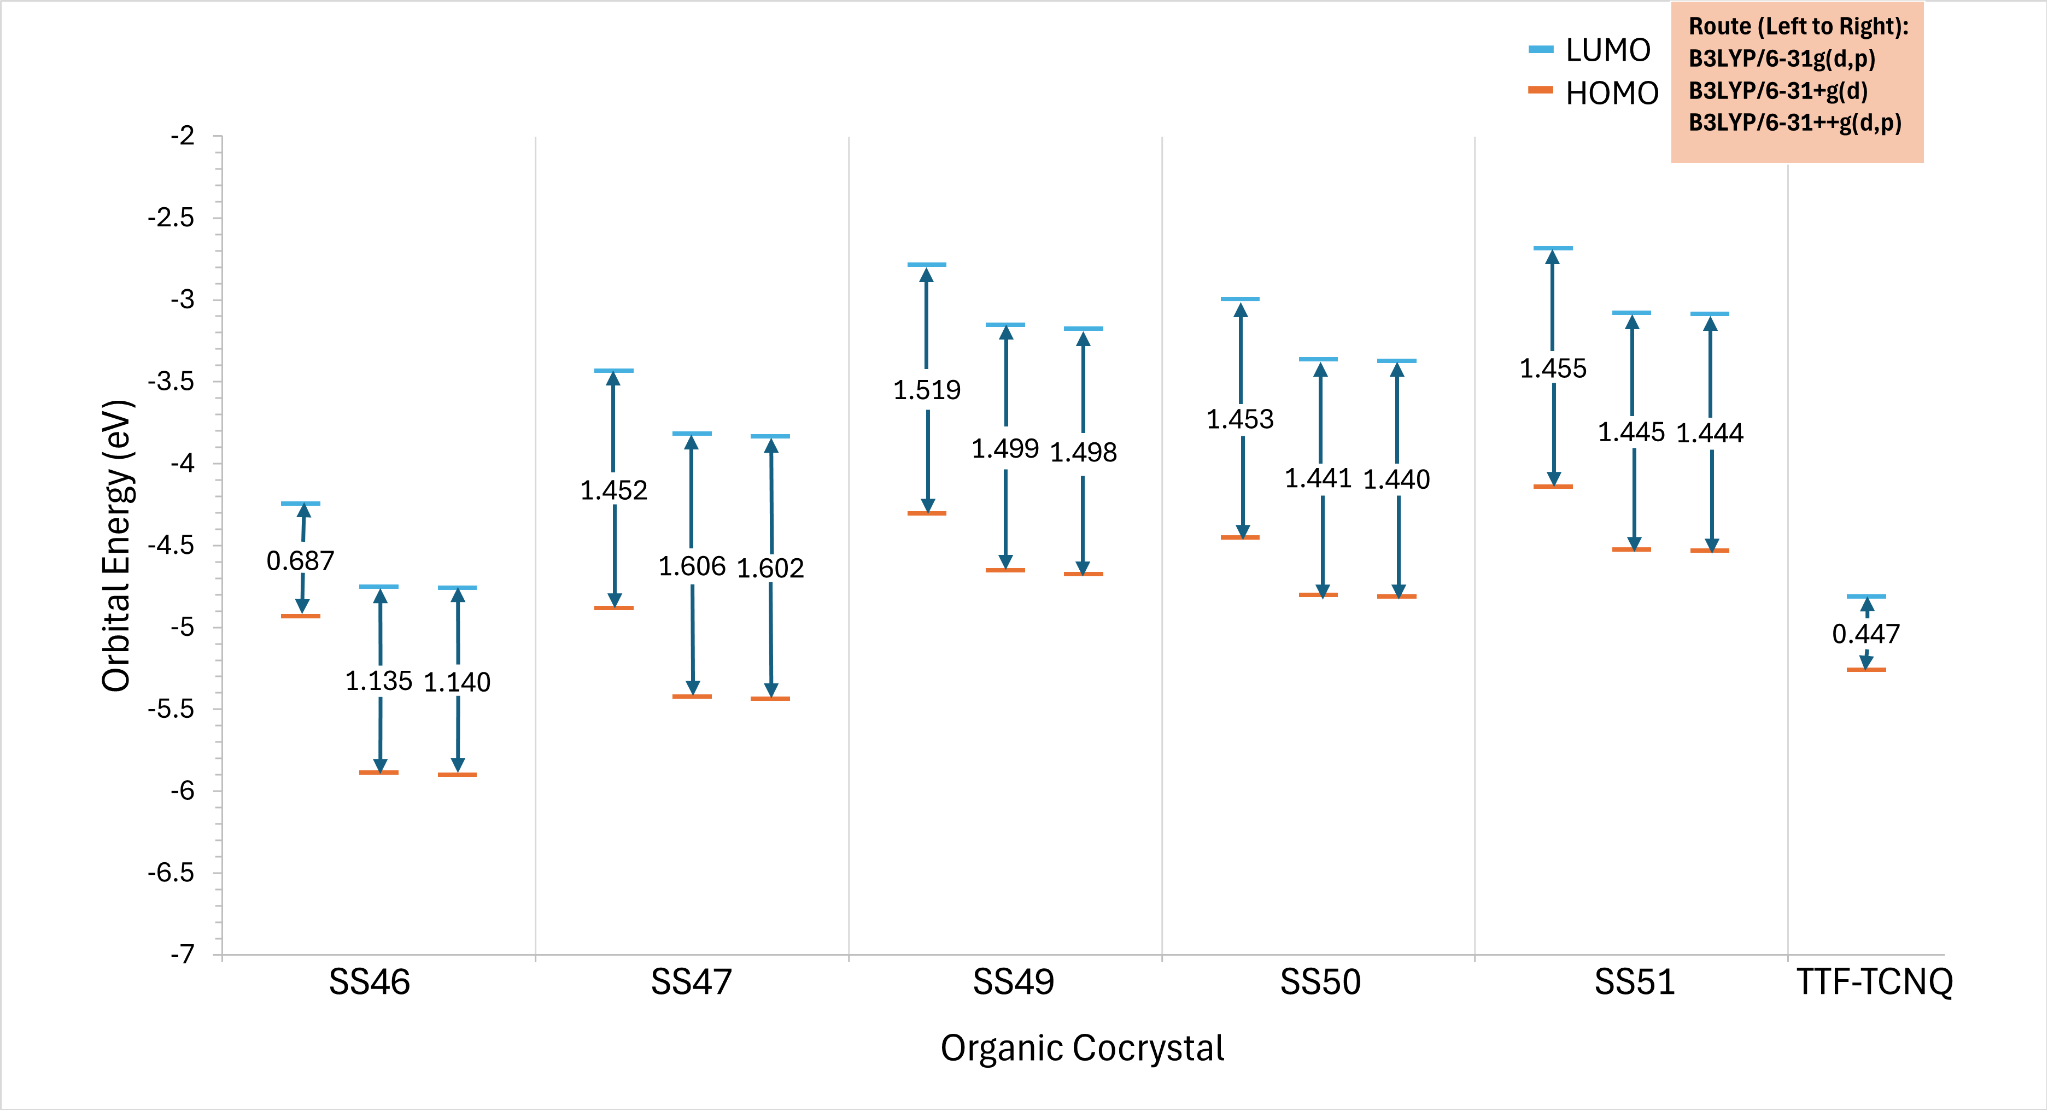
For isolated donor–acceptor pairs, we used Gaussian 16 with three B3LYP basis sets, checked optimized structures with frequency calculations, and collected the energies and locations of the frontier orbitals. We also calculated TTF–TCNQ under the same conditions as a reference.
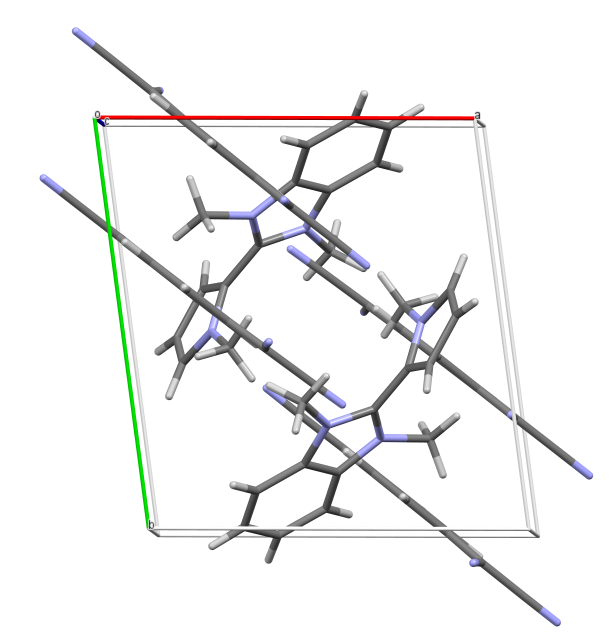
At the crystal scale, we used the X-ray-derived structural files in MedeA and VASP for periodic calculations, including electronic band structures and density of states. Keeping the two scales separate matters: a molecular HOMO–LUMO gap is not the same quantity as a crystal band gap.
From measured structure to calculated properties
X-ray diffraction supplied cell parameters and atomic coordinates. The computational workflow then evaluated electronic properties; the study did not measure electrical conductivity.
Molecular results
With the diffuse basis sets highlighted in the final poster, calculated HOMO–LUMO gaps for the new donor–acceptor pairs were broadly consistent at 1.1–1.5 eV. The TTF–TCNQ reference was approximately 0.4 eV under the same calculation conditions.
Orbital visualizations placed the frontier orbitals primarily on the TCNQ acceptor. This suggests limited donor–acceptor coupling for the isolated molecular pairs and motivates looking at the complete crystal.
Crystal results
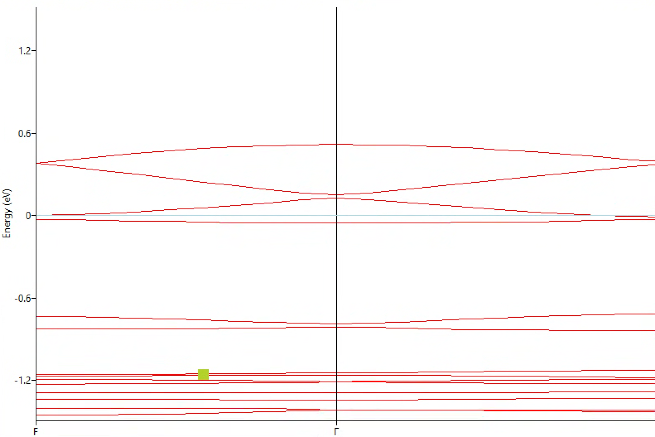
Initial periodic calculations put the band gaps of most candidates below 2 eV. SS49 had the narrowest calculated gap, 0.066 eV, making it a candidate for further investigation of charge transport. A narrow calculated gap suggests a possibility of higher conductivity; conductivity was not experimentally established here.
Next steps & resources
The calculations provided an initial comparison of the cocrystals and identified SS49 for closer study. Further analysis of the band structure, density of states, and charge transport was planned in the final poster. These results are computational predictions, not a measured device performance.
Poster authors: Yupeng Li, Kripa Shankar SR, Swagat K. Mohapatra, and Rakesh K. Behera. The study was carried out through NYU's Undergraduate Summer Research Program with support from NYU high-performance computing resources.